Electronic displacement effects play a crucial role in organic chemistry. They influence how molecules react and behave..
Electronic Displacement Effects in Organic Chemistry: Inductive, Resonance, Hyperconjugation & Electromeric Effects
Every predictable property of an organic molecule — its acidity, its stability as an intermediate, and the exact path a reaction takes — is, in fact, governed by how its electrons move. These movements, therefore, are called electronic displacement effects, and they are, ultimately, the single most important idea in physical organic chemistry. Moreover, this guide covers the four fundamental effects — inductive, resonance, hyperconjugation, and electromeric — along with complete +I/-I and +R/-R group lists, comparison tables, 10 fully solved problems.
Key Takeaways: Electronic Displacement Effects
- Electronic Effects Organic Chemistry are the four ways electrons shift within a molecule, and they control reactivity, acidity and stability.
- The inductive effect is permanent sigma-bond polarisation; +I groups donate, -I groups withdraw, and -NO2/-F/-CN are strong -I groups.
- Resonance delocalises pi electrons across a conjugated system; +R groups (-NH2, -OH) release, -R groups (-NO2, -CN) withdraw.
- Hyperconjugation (‘no-bond resonance‘) explains the carbocation stability order 3deg > 2deg > 1deg > methyl and alkene stability.
- The electromeric effect is temporary and reagent-induced, governing the regiochemistry of addition reactions.
- Master these four effects and you can predict reaction outcomes instead of memorising them.
What Are Electronic Displacement Effects?
Electronic displacement effects are the shifts of bonding or non-bonding electrons within a molecule caused by (1) an electronegativity difference between atoms, (2) the presence of a pi (pi) bond or conjugated system, (3) an adjacent charge, or (4) the approach of an attacking reagent. They are the conceptual foundation of physical organic chemistry, first organise into a unified framework by the British chemist Christopher Ingold, whose 1953 treatise Structure and Mechanism in Organic Chemistry grouped inductive, mesomeric (resonance) and electromeric effects under one theory that is still taught today.
Why Electrons Move
A covalent bond is rarely a perfectly equal sharing of electrons. When two atoms differ in electronegativity, the bonding pair is pulls toward the more electronegative atom, polarising the bond and creating a permanent dipole. That polarisation – and its knock-on effect on neighbouring bonds – is where all electronic displacement begins.
Impact on Reactivity, Acidity & Stability
These effects decide three things for every molecule: (1) which sites are electron-rich (nucleophilic) or electron-poor (electrophilic); (2) how stable charged intermediates such as carbocations and carbanions are; and (3) how acidic or basic a neighbouring functional group is. They are why acetic acid is acidic but ethanol is not, and why allyl and benzyl carbocations are unusually stable.
Inductive Effect (+I and -I) – Permanent Polarization
The inductive effect (I effect) is the permanent polarisation of a sigma (sigma) bond produces by an electronegativity difference between two adjacent atoms. Consider a C-Cl bond: chlorine is more electronegative than carbon, so the shared electron pair is drawn toward chlorine, giving it a partial negative charge (delta-) and carbon a partial positive charge (delta+). That dipole is not isolated – it pulls slightly on the next C-C bond, which pulls slightly on the one after that. The result is a chain of bond polarisation that travels down the molecule.
Crucially, the inductive effect fades rapidly with distance – after three to four carbon atoms it is practically negligible. It is also transmit only through sigma bonds and is never reversed. Because it is an intrinsic property of the molecule (not induces by an external attacking species), the inductive effect is defined as permanent – it is present whether or not a reaction is taking place.
The magnitude and order of the inductive effect are determine from Pauling’s electronegativity data and confirmed experimentally through pKa measurements. The logic is direct: an electron-withdrawing group near an acidic proton stabilises the conjugate base, lowering the pKa. Comparing the measures pKa of substituted acids therefore gives a quantitative ranking of -I strength, which is exactly how the standard group lists below were established.
+I Groups (Electron-Donating) – List with Examples
+I groups push electron density away from themselves toward the carbon chain, increasing electron density along the chain (and often stabilising a nearby positive charge). Alkyl groups are the classic +I donors: they are less electronegative than hydrogen relative to a carbon framework, so their effect is electron-releasing. The +I strength increases with chain length and branching because each additional C-C bond contributes to the forward push:
Alkyl groups (increasing +I): -C(CH3)3 (tert-butyl) > -CH(CH3)2 (isopropyl) > -CH2CH3 (ethyl) > -CH3 (methyl) > -H
Strong +I (negatively charged) groups: -O(-) (oxide), -COO(-) (carboxylate), -N(-)(C=O) (amide anion) – a full negative charge releases electrons far more strongly than any neutral group.
-I Groups (Electron-Withdrawing) – List with Examples
-I groups attract electron density away from the chain, decreasing electron density and stabilising a nearby negative charge. Their strength tracks electronegativity (and, for charged groups, a full positive charge). The standard -I strength order, established from pKa and dipole data, is:
-NO2 > -CN > -SO3H > -COOH > -F > -Cl > -Br > -I > -OH > -OR > -C6H5 > -H
Full list: -NO2 (nitro), -CN (cyano), -COOH (carboxyl), -CHO (formyl), -COR (acyl), -F, -Cl, -Br, -I (halogens), -OH (hydroxyl), -OR (alkoxy), -NH2 (amino), -C6H5 (phenyl).
Note the halogen flip: fluorine has the strongest -I effect among halogens (highest electronegativity), yet it is also a weak +R donor through its lone pair – a duality that appears again in electrophilic aromatic substitution.
3 Solved Problems: Electronic displacement effects
Arrange -CH3, -C2H5 and -C(CH3)3 in increasing +I strength. Answer: -CH3 < -C2H5 < -C(CH3)3. Branching increases the number of C-C bonds pushing electron density forward, so tert-butyl is the strongest +I donor here.
Which is more acidic – CH3COOH or ClCH2COOH? Chloroacetic acid is more acidic. The -I chlorine pulls electron density away from the O-H carboxyl, stabilising the chloroacetate anion and lowering the pKa.
Which is the stronger base – ethylamine or aniline? Ethylamine. The alkyl group is +I and donates density to nitrogen, whereas in aniline the lone pair is delocalises into the benzene ring, making it far less available for protonation.
Applications of Inductive Effect: Electronic Effects Organic Chemistry
The inductive effect is not an abstract concept – it predicts real, measurable chemistry. Each application below is verified against standard pKa and stability data from the literature.
Acidity of Carboxylic Acids
A -I substituent, in fact, stabilises the carboxylate ion (RCOO-) by pulling electron density away from the negatively charged oxygen, thereby spreading and stabilising the negative charge. Consequently, this shifts the deprotonation equilibrium toward products and increases acidity. Therefore, the stronger the -I effect, the lower the pKa. Hence, the measurable acidity order:
Cl3CCOOH (trichloroacetic, pKa ~0.65) > Cl2CHCOOH (dichloroacetic, pKa ~1.29) > ClCH2COOH (chloroacetic, pKa ~2.86) > CH3COOH (acetic, pKa ~4.76)
Each additional -I chlorine lowers the pKa further because it draws even more density off the carboxylate. Distance matters too: a chlorine on the beta carbon has far less effect than one on the alpha carbon, because the inductive effect decays rapidly along the chain.
Stability of Carbocations: Electronic Displacement Effects
+I alkyl groups attached to a positively charged carbon donate electron density and disperse the positive charge, stabilising the carbocation – the ion is lower in energy when its charge is spread out. The usual stability order is tertiary (3deg) > secondary (2deg) > primary (1deg) > methyl, because each additional alkyl group provides another +I source. This ordering is exactly why SN1 reactions proceed fastest through tertiary centres and cannot proceed at all through methyl centres.
Reactivity of Alkyl Halides
Alkyl halides show the two effects competing. In an SN1 reaction the rate-determining step forms a carbocation, so +I groups that stabilise that cation accelerate the reaction – tertiary alkyl halides react fastest via SN1. In SN2, backside attack on the electrophilic carbon is rate-limiting and steric hindrance dominates – primary halides react fastest, and tertiary halides barely react at all. Being able to predict SN1 vs SN2 from +I effects and sterics is a core exam skill; see our reaction types guide for the complete mechanism.
Basicity of Amines
+I alkyl groups push electron density onto the nitrogen lone pair, making alkyl amines more basic than ammonia in the gas phase – electron density on nitrogen is what makes it able to accept a proton. The gas-phase order is (CH3)3N > (CH3)2NH > CH3NH2 > NH3. In water the order is modified by solvation effects (hydrogen bonding to the ammonium ion), a reminder that the inductive effect is one factor among several when a solvent is present. Electron-withdrawing groups near nitrogen always reduce basicity by draining density from the lone pair.
Resonance Effect (+R and -R) – Delocalization of pi Electrons
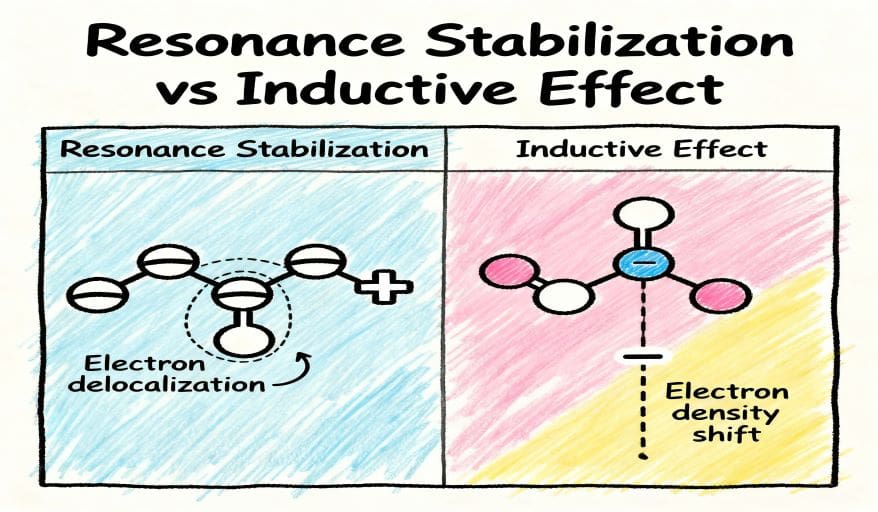
The resonance (or mesomeric) effect, in fact, is the delocalisation of pi electrons — or of a lone pair — across a conjugated system of parallel p orbitals. Moreover, resonance theory was introduced by Linus Pauling, who demonstrated that a single Lewis structure is often inadequate for describing a molecule. Instead, the true structure is a hybrid of several contributing ‘canonical’ forms. Consequently, the real molecule is lower in energy (more stable) than any single canonical form. Therefore, this lowering of energy is called resonance stabilisation or resonance energy.
For resonance to occur, a molecule must satisfy three conditions: (1) it must have a conjugated pi system (alternating single and double bonds) or a lone pair on an atom adjacent to a pi bond; (2) the atoms involved must be coplanar or nearly coplanar so their p orbitals can overlap side-on; and (3) every contributing form must be a valid Lewis structure. If any of these fails, delocalisation is blocks and the effect is lost.
+R Groups (Electron-Releasing by Resonance)
+R (or +M) groups donate pi electron density into the system, usually through a lone pair. Typical +R order:
-O(-) > -NH2 > -NHR > -OH > -OR > -NHCOR > -OCOR > -C6H5
Halogens (-F, -Cl, -Br, -I) show a weak +R effect through their lone pairs even though their -I effect dominates overall.
-R Groups (Electron-Withdrawing by Resonance)
-R (or -M) groups withdraw pi density, usually through a C=O, C=N or polarised multiple bond. Typical -R order:
-NO2 > -CN > -CHO > -COR > -COOH > -COOR > -SO3H
Drawing Resonance Structures – Rules & 5 Examples
Rules: (1) first, only electrons move — never atoms or nuclei; (2) moreover, every canonical form must be a valid Lewis structure; (3) additionally, keep the same total number of unpaired electrons; (4) furthermore, use curved arrows to show electron flow; (5) finally, the resonance hybrid — not any single form — is the real, most stable structure.
Nitrate ion (NO3-) – the negative charge is shared equally over three oxygen atoms in the hybrid.
Benzene – two Kekule structures combine into a symmetric hybrid with partial double-bond character, explaining its unusual stability.
Carboxylate ion (RCOO-) – the negative charge is delocalised over two equivalent oxygens, stabilising it and driving acidity.
Nitrobenzene – the -NO2 group withdraws from the ring, placing positive charge at ortho and para positions.
Phenol – the oxygen lone pair delocalises into the ring, placing negative charge at ortho and para carbons.
Applications of Resonance Effect: Electronic Effects Organic Chemistry
Acidity of Phenols vs Alcohols
Phenol (pKa around 10) is about six orders of magnitude more acidic than ethanol (pKa around 16). The reason is resonance: the phenoxide ion delocalises its negative charge across the aromatic ring – the charge appears at the oxygen and at the ortho and para carbons in the resonance hybrid. An alkoxide ion (RO-) has no such delocalisation and holds the negative charge tightly on a single oxygen, so it is far less willing to form. The resonance energy of the phenoxide ion is what makes phenol acidic.
Stability of Benzyl and Allyl Carbocations
Benzyl and allyl carbocations are stabilised by resonance, which spreads the positive charge over multiple carbons rather than pinning it on one. In the benzyl cation (C6H5CH2(+)), the positive charge is shared by the benzylic carbon and the ortho and para ring carbons; in the allyl cation (CH2=CH-CH2(+)), it is shared equally by the two terminal carbons. This is why benzyl and allyl halides undergo SN1 reactions far more readily than ordinary alkyl halides – the intermediate cation is unusually stable, so it forms faster.
Directing Effects in Electrophilic Aromatic Substitution: Electronic displacement effects
+R groups such as -OH and -NH2 activate the ring and direct incoming electrophiles to the ortho and para positions, because those are the positions where the resonance donor places extra electron density. -R groups such as -NO2 and -CHO deactivate the ring and direct meta, because they withdraw density from ortho and para and leave meta the least disfavoured. This is a direct application of the resonance effect, and it is where many Class 11 students first see the concept pay off.
Directing Effects in Electrophilic Aromatic Substitution (Preview)
+R groups such as -OH and -NH2 activate the ring and direct incoming electrophiles to the ortho and para positions; -R groups such as -NO2 and -CHO deactivate the ring and direct meta. Coming soon: how resonance and electronic effects control electrophilic aromatic substitution – a complete guide to aromatic chemistry.
Hyperconjugation – No-Bond Resonance
Hyperconjugation is the delocalising interaction of sigma-bond electrons (usually a C-H bond, sometimes a C-C bond) with an adjacent unsaturated centre – an empty p orbital, a pi bond, or a radical. It is often called ‘no-bond resonance’ because it involves sigma electrons rather than pi electrons, and it is historically known as the Baker-Nathan effect.
It requires at least one alpha-hydrogen: a hydrogen attached to the carbon directly adjacent to the unsaturated centre. In the methyl carbocation CH3(+), for example, each of the three C-H sigma bonds can donate density into the empty p orbital, so the positive charge is delocalises over three hydrogens as well as the carbon – the C-H bond temporarily resembles a pi-type interaction. Every alpha C-H bond is a potential hyperconjugative donor, which is why counting alpha-hydrogens is the quickest way to compare stability.
Hyperconjugation in Alkenes (Stability Order)
The more alpha-hydrogens an alkene has, the more hyperconjugative structures it can form, and the more stable the alkene. Each alpha C-H sigma bond can donate into the adjacent pi* orbital, so stability increases with the degree of alkyl substitution on the double bond:
CH2=CH2 (ethene, no alkyl groups) < RCH=CH2 (monosubstituted) < RCH=CHR (disubstituted) < R2C=CR2 (tetrasubstituted)
This is why 2-butene is more stable than 1-butene, and why tetrasubstituted alkenes are the most stable of all – more substituents mean more sigma donors into the pi system.
Hyperconjugation in Carbocations
Hyperconjugation, in fact, explains the classic carbocation stability order 3deg > 2deg > 1deg > methyl. In a tertiary carbocation, for example, nine alpha C-H bonds are available to delocalise the positive charge into its empty p orbital. Similarly, a secondary has six, a primary has three, and the methyl cation has none. Therefore, the more no-bond resonance structures, the more the positive charge is dispersed, and consequently, the lower the energy of the cation.
This is not just a ranking to memorise – it is the mechanistic reason behind real observations: tertiary haloalkanes favour SN1 because their carbocation forms easily, tertiary alcohols dehydrate readily, and Markovnikov’s rule follows carbonation stability. Carbocation stability from hyperconjugation explains why tertiary haloalkanes favour SN1 – see our haloalkanes and haloarenes guide.
3 Solved Problems: Electronic displacement effects
Arrange by stability: (CH3)3C(+), (CH3)2CH(+), CH3CH2(+), CH3(+).
Answer: (CH3)3C(+) > (CH3)2CH(+) > CH3CH2(+) > CH3(+). More alkyl groups supply more alpha-hydrogens for hyperconjugation and a stronger +I effect.
Which alkene is more stable – CH3CH=CH2 or CH3CH=CHCH3? 2-butene (CH3CH=CHCH3) – the more substituted double bond offers more hyperconjugative stabilisation.
Why does propene have a small net dipole moment with the methyl end positive? The methyl group is a weak +I and hyperconjugative donor, pushing electron density toward the double bond.
Electromeric Effect – Temporary Displacement
The electromeric effect (E effect) is a temporary, complete transfer of a shared pi-electron pair to one of the atoms of a multiple bond in the presence of an attacking reagent, and it vanishes the instant the reagent is removed. This is the single most important contrast in electronic-displacement theory: the inductive and resonance effects are permanent, intrinsic features of the molecule, whereas the electromeric effect exists only in the transition state, while an external reagent is actually attacking.
Think of it, instead, as a reaction-time phenomenon, not a molecular property. In the transition state, the pi pair can, in fact, shift completely (not partially, as in the inductive effect) to one atom. As a result, this creates a momentary charge separation that drives the reaction to the product. Once the product forms, however, the electromeric effect has vanished. Therefore, it leaves no trace in the product molecule.
+E and -E Effects
+E effect – the pi electron pair shifts completely toward the attacking electrophile, creating a negative centre where the electrophile attacks. Example: H(+) approaching an alkene polarises the C=C bond toward the incoming H(+).
-E effect – the pi electron pair shifts completely away from the attacking nucleophile, creating a positive centre where the nucleophile attacks. Example: CN(-) approaching a carbonyl carbon pushes the C=O pi pair up onto oxygen, exposing the electrophilic carbon.
Role in Addition Reactions
The electromeric effect explains the momentary polarisation of the transition state in alkene and carbonyl addition reactions. It determines the regiochemical outcome – which carbon the electrophile adds to first – and is central to Markovnikov-type addition patterns. In the electrophilic addition of HX to propene, the +E effect directs H(+) to the terminal carbon, forming the more stable secondary carbocation that then captures the halide.
10 Practice Problems with Step-by-Step Solutions
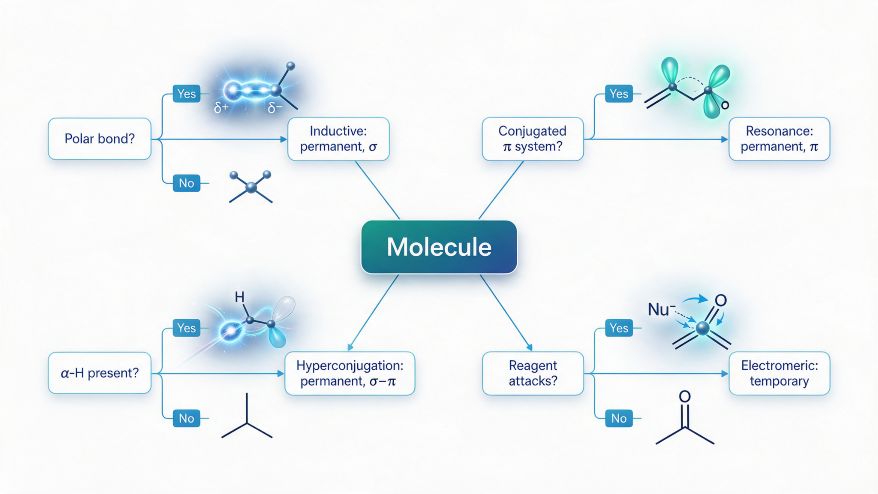
These problems of Electronic displacement effects:
- Predict the order of +I effect for: -CH3, -CH2CH3, -CH(CH3)2.
Solution: -CH2CH3 > -CH(CH3)2 vs -CH3? Series by increasing +I: -CH3 < -CH2CH3 < -CH(CH3)2. - Which carboxylic acid is strongest – acetic, chloroacetic, or trichloroacetic?
Solution: trichloroacetic > chloroacetic > acetic; more -I chlorine atoms stabilise the carboxylate more. - Arrange Cl, F, Br by -I strength.
Solution: -F > -Cl > -Br (electronegativity decreases down the group, so -I decreases). - Which is more stable – (CH3)3C(+) or CH3CH2(+)?
Solution: (CH3)3C(+); three +I methyl groups disperse the positive charge. - Which amine is more basic – methylamine or trimethylamine?
Solution: in the gas phase, trimethylamine (greater +I); in water the order flips due to solvation – always specify the phase. - Why is a carboxylate ion more stable than an alkoxide ion?
Solution: the carboxylate delocalises its negative charge over two equivalent oxygens; an alkoxide cannot. - Draw two significant canonical forms of the nitro group attached to a benzene ring.
Solution: -NO2 withdraws pi density, placing positive charge at ortho/para carbons in the resonance hybrid. - Why is phenol acidic while ethanol is essentially neutral?
Solution: the phenoxide ion is resonance-stabilised; the ethoxide ion is not, raising phenol’s acidity by about 10^6. - Explain the stability order 2deg carbocation > 1deg carbocation.
Solution: the 2deg cation has six alpha C-H sigma bonds vs three for the 1deg cation, giving more no-bond resonance structures. - Which alkene is more stable – 1-butene or 2-butene?
Solution: 2-butene; it is more substituted, hence more hyperconjugative stabilisation.
Comparison Table: Electronic Effects Organic Chemistry
| Effect | Type / Bond | Permanent or Temporary | Transmission | Direction | Key Examples |
| Inductive | sigma bond polarisation | Permanent | Along sigma bonds, fades in 3-4 atoms | +I donates; -I withdraws | -CH3 (+I); -NO2, -F, -CN (-I) |
| Resonance (Mesomeric) | pi electrons / lone pair in conjugation | Permanent | Across conjugated pi system | +R releases; -R withdraws | -NH2, -OH (+R); -NO2, -CN, -CHO (-R) |
| Hyperconjugation | sigma C-H with adjacent pi / empty p | Permanent (no-bond resonance) | Adjacent sigma C-H to unsaturated centre | Donating (like +I) | Stability of 2deg/3deg carbocations and substituted alkenes |
| Electromeric | pi electron pair, reagent-induced | Temporary (only during attack) | Instantaneous in transition state | +E toward electrophile; -E away from nucleophile | H+ addition to alkene (+E); CN- attack on carbonyl (-E) |
Frequently Asked Questions: Electronic Effects Organic Chemistry
What are electronic displacement effects in organic chemistry?
They are, in fact, the shifts of bonding and non-bonding electrons caused by electronegativity differences, conjugation, adjacent charge, or an attacking reagent. Moreover, the four main types are inductive, resonance, hyperconjugation, and electromeric effects. Therefore, these effects help explain how electron distribution changes within organic molecules.
What is the difference between the inductive and electromeric effects?
The inductive effect is a permanent polarisation of sigma bonds present even without a reaction. The electromeric effect is a temporary, complete transfer of a pi-electron pair that occurs only while a reagent is attacking, and disappears when the reagent is removed.
What is hyperconjugation with an example?
Hyperconjugation, in fact, is the delocalisation of sigma C-H electrons into an adjacent empty p orbital or pi bond. For example, the methyl groups in (CH3)3C(+) donate through their C-H bonds. As a result, this makes the tertiary carbocation the most stable.
What is the difference between +I and -I effect?
+I groups (e.g. alkyl groups) push electron density away from themselves and stabilise positive charge. -I groups (e.g. -NO2, -F, -CN) pull electron density toward themselves and stabilise negative charge.
How does resonance affect acidity?
Resonance stabilises the conjugate base by spreading its negative charge, which shifts the equilibrium toward deprotonation and increases acidity. This is why phenol (pKa ~10) is far more acidic than ethanol (pKa ~16).
References:
- D. Simatos, I. E. Jacobs, I. Dobryden, M. Nguyen, A. Savva, D. Venkateshvaran, M. Nikolka, J. Charmet, L. J. Spalek, M. Gicevičius, Y. Zhang, G. Schweicher, D. J. Howe, S. Ursel, J. Armitage, I. B. Dimov, U. Kraft, W. Zhang, M. Alsufyani, I. McCulloch, R. M. Owens, P. M. Claesson, T. P. J. Knowles, H. Sirringhaus, Effects of Processing-Induced Contamination on Organic Electronic Devices. Small Methods 2023, 7, 2300476. https://doi.org/10.1002/smtd.202300476
This article was written by Juveriya Khan and reviewed for editorial accuracy by our editorial team. It has not yet undergone independent review by a professional.
- Author
- Latest Posts
I am an inspirational writer and emerging science blogger with a deep-rooted passion for understanding and sharing the wonders of the world around us. My journey began with a Bachelor of Science in Chemistry from AKI’s Poona College of Arts, Commerce and Science, where I developed not only a strong foundation in scientific principles but also a mindset that values curiosity, observation, and critical thinking. Studying chemistry has taught me to approach problems with logic, to look beyond the obvious, and to see patterns where others might only see complexity. These experiences have shaped the way I perceive life and influence the way I express my thoughts, both scientifically and creatively.
I have always believed that knowledge is only as powerful as the way it is shared. This belief has guided me to create content that bridges the gap between science and everyday life. I aim to simplify complex scientific ideas so they become approachable, relatable, and engaging for everyone—whether someone has a formal background in science or is simply curious about the world. Science, in my view, is not confined to textbooks and laboratories; it is a way of thinking, understanding, and connecting with the world around us. By making science accessible, I hope to inspire curiosity, ignite learning, and encourage people to see the beauty and relevance of scientific discovery in their daily lives.
Alongside my love for science, I am deeply passionate about motivational writing. I believe words carry the power to transform perspectives, encourage self-reflection, and inspire action. By combining analytical thinking with creativity, I create content that is intellectually stimulating yet emotionally resonant. Whether I am breaking down a chemical concept, exploring a scientific phenomenon, or crafting motivational narratives, my goal is to communicate ideas in a way that is clear, meaningful, and impactful.
Through my work, I strive to empower my readers – not just by helping them understand the science around them, but by inspiring them to grow, learn, and realize their full potential. Every article, post, or story I write reflects my commitment to turning knowledge into inspiration. I want to spark curiosity, encourage self-improvement, and show that learning and personal growth are deeply connected. For me, writing is not just a craft; it is a way to touch minds, connect hearts, and make ideas come alive in a way that truly resonates.